Collecting data
Collecting data
Accession list of SARS-like viruses:
In[]:=
viruses={"AY274119.3","AY278488.2","AY278491.2","AY278554.2","AY278741.1","AY282752.2","AY283794.1","AY283795.1","AY283796.1","AY283797.1","AY283798.2","AY291451.1","AY304488.1","AY390556.1","AY394985.1","AY394996.1","AY395003.1","AY502927.1","AY559093.1","EU371559.1","EU371562.1","EU371564.1","GU553365.1","KF294455.1","KJ473816.1","KY417144.1","KY417145.1","KY967357.1","KY983587.1","MF542265.1","MG772933.1","MG772934.1","MK211376.1","NC_002645.1","NC_004718.3","NC_005831.2","NC_019843.3","NC_038294.1","MN996532","KF367457"};
One can use the function ImportFASTA, from Wolfram Function Repository, to Import FASTA data from the NCBI using an NCBI Reference Sequence. To save time, I have done it and gave it an iconized form.
In[]:=
virusNameSeq=(*Map[ResourceFunction["ImportFASTA"],viruses]*);
Take 1st element of from all sublist of virusNameSeq, flatten the result, and return the substrings from the first space to “,”:
In[]:=
names=Flatten[StringCases[virusNameSeq[[All,1]]//Flatten," "~~x__~~","x]]
Out[]=
{Severe acute respiratory syndrome-related coronavirus isolate Tor2,SARS coronavirus BJ01,SARS coronavirus HKU-39849,SARS coronavirus CUHK-W1,SARS coronavirus Urbani,SARS coronavirus CUHK-Su10,SARS coronavirus Sin2500,SARS coronavirus Sin2677,SARS coronavirus Sin2679,SARS coronavirus Sin2748,SARS coronavirus Sin2774,SARS coronavirus TW1,SARS coronavirus SZ16,SARS coronavirus GZ02,SARS coronavirus HSZ-Bb,SARS coronavirus ZS-B,SARS coronavirus ZS-C,SARS coronavirus TW4,SARS coronavirus Sin845,SARS coronavirus ZJ02,SARS coronavirus BJ182-4,SARS coronavirus BJ182-12,SARS coronavirus HKU-39849 isolate TCVSP-HARROD-00003,SARS-related bat coronavirus isolate Anlong-111 orf1ab polyprotein and orf1a polyprotein genes,BtRs-BetaCoV/YN2013,Bat SARS-like coronavirus isolate Rs4084,Bat SARS-like coronavirus isolate Rf4092,Human coronavirus 229E strain SC2872,Human coronavirus 229E strain SC3112,Human coronavirus 229E strain 229E/Haiti-1/2016,Bat SARS-like coronavirus isolate bat-SL-CoVZC45,Bat SARS-like coronavirus isolate bat-SL-CoVZXC21,Coronavirus BtRs-BetaCoV/YN2018B,Human coronavirus 229E,SARS coronavirus,Human Coronavirus NL63,Middle East respiratory syndrome coronavirus,Betacoronavirus England 1,Bat coronavirus isolate RaTG13,Bat SARS-like coronavirus WIV1}
Replacement rule to change U by T and keep only A, C, G and T:
In[]:=
srules={"U""T",Except[Characters["ACGT"]]""};
Give an association with viruses accession as keys and sequences as values, then replace the sequences using srules:
In[]:=
dataSARSCoV=StringReplace[srules]/@AssociationThread[viruses,Flatten[virusNameSeq[[All,2]]]];
Take COVID-19 data from WDR, then select those whole GenBankTitle contains “complete genome””
In[]:=
dataWDR=ResourceData["Genetic Sequences for Novel Coronavirus 2019-nCoV from Wuhan, China"][Select[StringContainsQ[#GenBankTitle,"complete genome"]&]];
From previous imported data, take only Accession and Sequence columns, make it as association with accession as keys and sequence as values, finally replace the sequences using srules:
In[]:=
dataCOVID19=StringReplace[srules]/@<|Rule@@@Normal[Values[dataWDR[All,{"Accession","Sequence"}]]]|>;
Generate an association from SARS-like and COVID-19 viruses with accessions as keys and corresponding names as values:
In[]:=
accessionNameAssoc=Join[AssociationThread[viruses(Style[#,Bold,Italic]&/@names)],AssociationThread[Keys[dataCOVID19]StringSplit[Normal[dataWDR[All,"GenBankTitle"]],","][[All,1]]]];
Z-curve
Z-curve
For a given genetic sequence of length N, let’s denote the number of occurrence of A, C, G, and T bases at the n-th subsequence by , , , and . Then, the corresponding 3D coordinate for the n-th subsequence is obtained by the following transformation:
A
n
C
n
G
n
T
n
Out[]//TraditionalForm=
x n |
y n |
z n |
1 | -1 | 1 | -1 |
1 | 1 | -1 | -1 |
1 | -1 | -1 | 1 |
A n |
C n |
G n |
T n |
For example, for a sequence such as AACGT, we have: , , , , and , and the corresponding Z-curve would be {{0,0,0},{1,1,1},{2,2,2},{1,3,1},{2,2,0},{1,1,1}}.
{=1,=0,=0,=0}
A
1
C
1
G
1
T
1
{=2,=0,=0,=0}
A
2
C
2
G
2
T
2
{=2,=1,=0,=0}
A
3
C
3
G
3
T
3
{=2,=1,=1,=0}
A
4
C
4
G
4
T
1
{=2,=1,=1,=1}
A
5
C
5
G
5
T
5
Note that since =(+)-(+)it is the different between the Purine bases (R) and Pyrimidine ones (Y). Therefore, it is a measure of RY disparity. Likewise, quantifies the Amino (M) and Keto (K) difference, called as MK-disparity and the WS-disparity (W for Weak and S for Strong Hydrogen bonds).
x
n
A
n
G
n
C
n
T
n
y
n
z
n
Generate Z-curve by cumulative sums of four matrices assigned to A, C, G and T:
In[]:=
ZcurveGenerator[st_]:={{1,-1,1,-1},{1,1,-1,-1},{1,-1,-1,1}}.#&/@FoldList[#1+Switch[#2,"A",{1,0,0,0},"C",{0,1,0,0},"G",{0,0,1,0},"T",{0,0,0,1}]&,{0,0,0,0},Characters[st]]
Generate Z-curves of COVID-19 viruses (the result is an association in the form of accession->Z-curve):
In[]:=
zCurveCOVID19=ZcurveGenerator/@dataCOVID19;
Generate Z-curves of SARS-like viruses (the result is an association in the form of accession->Z-curve):
In[]:=
zCurveSARS=ZcurveGenerator/@dataSARSCoV;
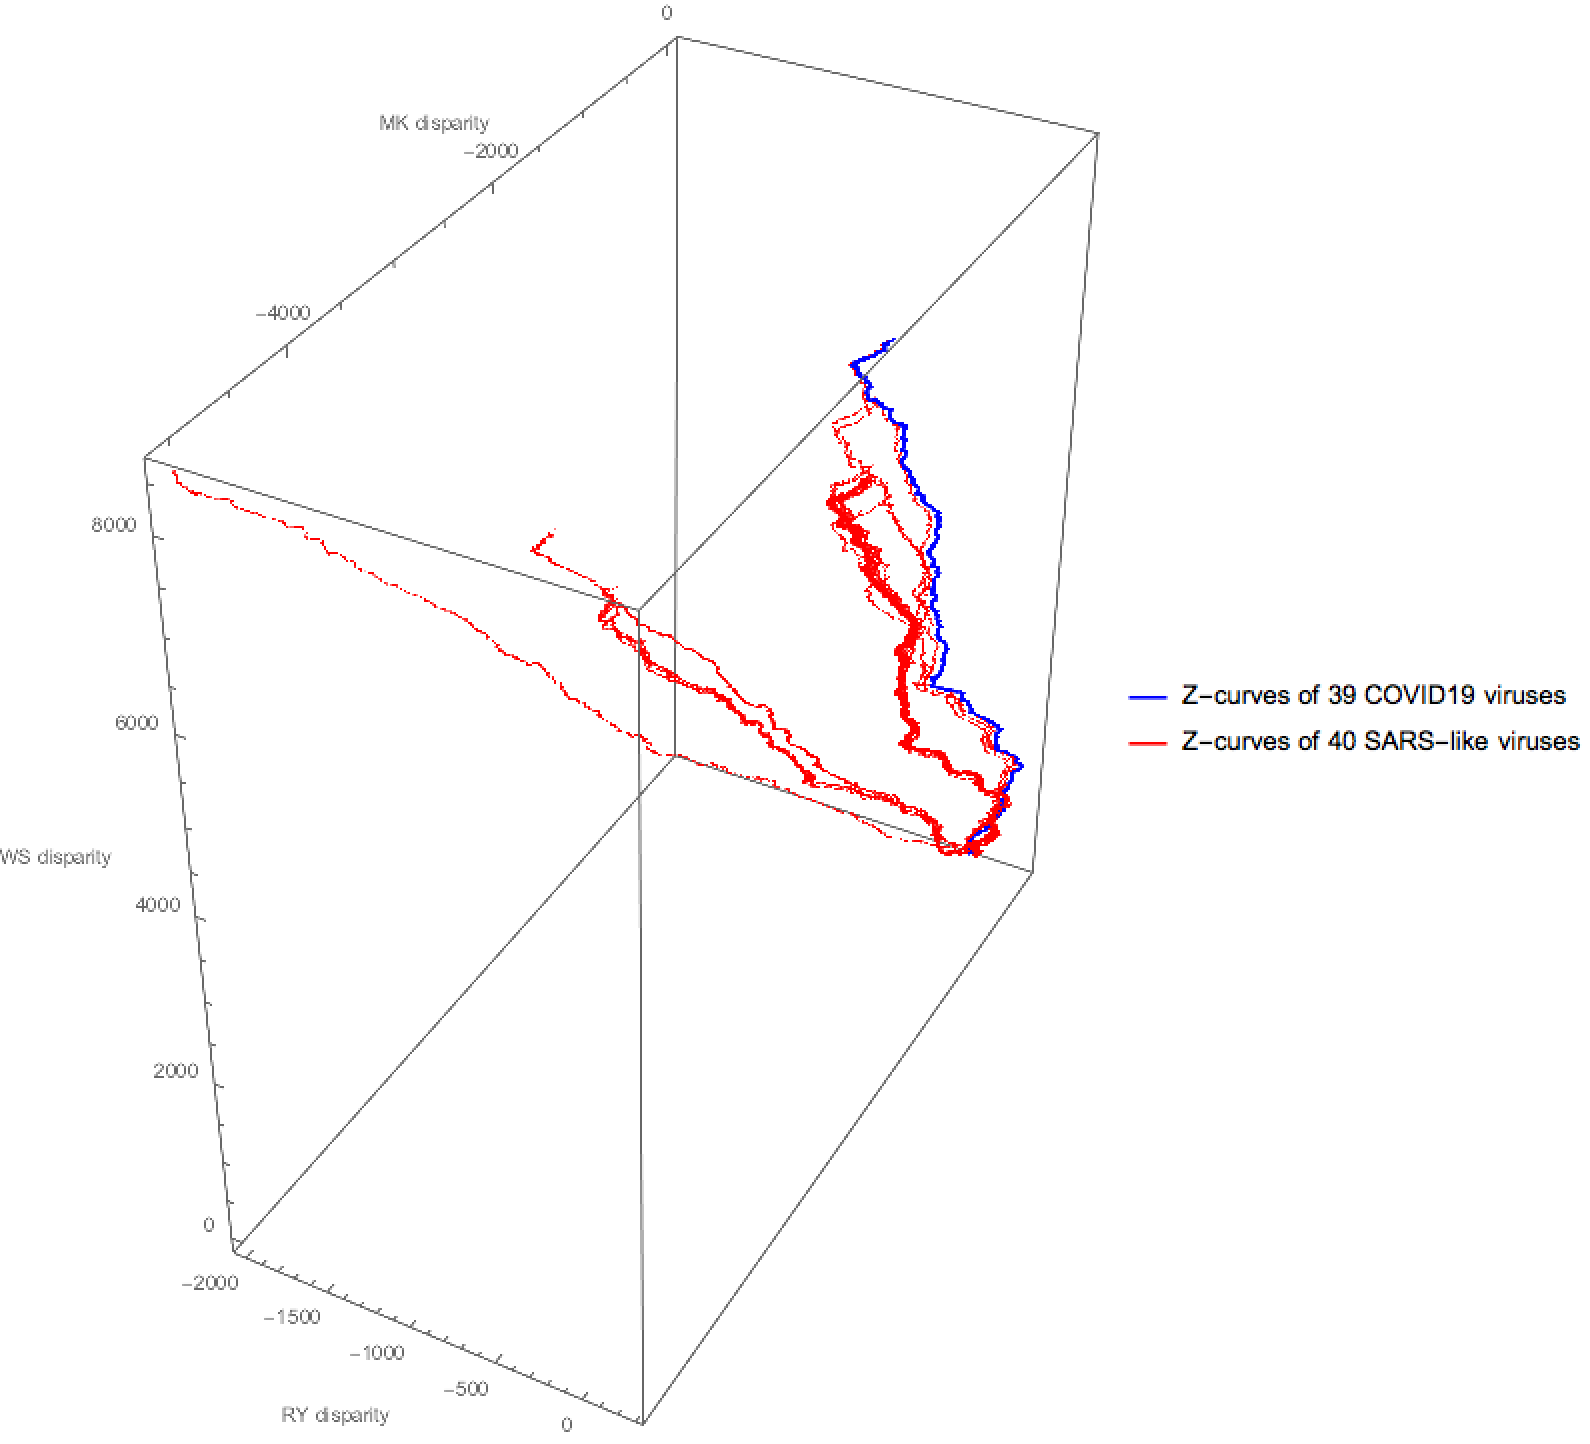
Represent 3D image of Z-curve where the red ones belongs to SARS-like viruses and blue ones of COVID-19:
Legended[Graphics3D[{Blue,Values[Line/@zCurveCOVID19],Red,Values[Line/@zCurveSARS]},AxesTrue,ImageSizeLarge,BoxRatios{2,4,4},AxesLabel{"RY disparity","MK disparity","WS disparity"}],LineLegend[{Blue,Red},{"Z-curves of 39 COVID19 viruses","Z-curves of 40 SARS-like viruses"}]]
Out[]=
Find the shortest sequence and denote its length by “min”:
In[]:=
min=Min[Min[Length/@zCurveCOVID19],Min[Length/@zCurveSARS]];
Trim the Z-curves up to the length of shortest sequence:
In[]:=
trimmedZCurveCOVID19=Take[#,min]&/@zCurveCOVID19;trimmedzCurveSARS=Take[#,min]&/@zCurveSARS;
Clustering and classification based on Z-curves
Clustering and classification based on Z-curves
Find the 3 nearest Z-curves of SARS-like viruses to the Z-curve of each COVID-19 viruses:
In[]:=
closest=Nearest[trimmedzCurveSARS,Values[trimmedZCurveCOVID19],3];
In[]:=
Grid[Prepend[Flatten/@Transpose[{Keys[trimmedZCurveCOVID19],closest}],{Style["COVID-19 Accession",Bold],Style["3 nearest SARS-like viruses",Bold],SpanFromLeft,SpanFromLeft}],FrameAll]
Out[]=
COVID-19 Accession | 3 nearest SARS-like viruses | ||
MN908947 | MN996532 | MG772934.1 | MG772933.1 |
NC_045512 | MN996532 | MG772934.1 | MG772933.1 |
MN938384 | MN996532 | MG772934.1 | MG772933.1 |
MN975262 | MN996532 | MG772934.1 | MG772933.1 |
MN985325 | MN996532 | MG772934.1 | MG772933.1 |
MN988713 | MN996532 | MG772934.1 | MG772933.1 |
MN988668 | MN996532 | MG772934.1 | MG772933.1 |
MN988669 | MN996532 | MG772934.1 | MG772933.1 |
MN994467 | MN996532 | MG772934.1 | MG772933.1 |
MN994468 | MN996532 | MG772934.1 | MG772933.1 |
MN997409 | MN996532 | MG772934.1 | MG772933.1 |
MN996527 | MN996532 | MG772934.1 | MG772933.1 |
MN996528 | MN996532 | MG772934.1 | MG772933.1 |
MN996529 | MN996532 | MG772934.1 | MG772933.1 |
MN996530 | MN996532 | MG772934.1 | MG772933.1 |
MN996531 | MN996532 | MG772934.1 | MG772933.1 |
MT007544 | MN996532 | MG772934.1 | MG772933.1 |
MT019529 | MN996532 | MG772934.1 | MG772933.1 |
MT019530 | MN996532 | MG772934.1 | MG772933.1 |
MT019531 | MN996532 | MG772934.1 | MG772933.1 |
MT019532 | MN996532 | MG772934.1 | MG772933.1 |
MT019533 | MN996532 | MG772934.1 | MG772933.1 |
MT020880 | MN996532 | MG772934.1 | MG772933.1 |
MT020881 | MN996532 | MG772934.1 | MG772933.1 |
LC521925 | MN996532 | MG772934.1 | MG772933.1 |
MT027062 | MN996532 | MG772934.1 | MG772933.1 |
MT027063 | MN996532 | MG772934.1 | MG772933.1 |
MT027064 | MN996532 | MG772934.1 | MG772933.1 |
LC522972 | MN996532 | MG772934.1 | MG772933.1 |
LC522973 | MN996532 | MG772934.1 | MG772933.1 |
LC522974 | MN996532 | MG772934.1 | MG772933.1 |
LC522975 | MN996532 | MG772934.1 | MG772933.1 |
MT039873 | MN996532 | MG772934.1 | MG772933.1 |
MT039887 | MN996532 | MG772934.1 | MG772933.1 |
MT039888 | MN996532 | MG772934.1 | MG772933.1 |
MT039890 | MN996532 | MG772934.1 | MG772933.1 |
MT044257 | MN996532 | MG772934.1 | MG772933.1 |
MT044258 | MN996532 | MG772934.1 | MG772933.1 |
MT049951 | MN996532 | MG772934.1 | MG772933.1 |
As suggested by the Z-curve visualization, three nearest SARS-like viruses are the same for all COVID-19 viruses.
Delete duplicates from the previous result and return their GenBank titles:
Our analysis shows that among 40 SARS-like viruses that we study here, Bat coronavirus isolate RaTG13, Bat SARS-like coronavirus isolate bat-SL-CoVZXC21, and Bat SARS-like coronavirus isolate bat-SL-CoVZC45 have closest genome similarity to COVID-19 viruses, providing evidence for the Bat origin of COVID-19.
Let us do some hierarchical clustering, too.
Concatenate Z-curves of COVID-19 and SARS-like viruses:
Create a numeric array of the type double-precision real our of each Z-curve:
Cluster the viruses based on their Z-curves using single-linkage clustering algorithm:
Assign different color to each cluster, and represent 3D images of corresponding Z-curves:
Project clusters into 3D using t-distributed stochastic neighbor embedding algorithm:
Constructs a dendrogram from the hierarchical clustering of the Z-curves (note the coloring is the same as the one for clusters and those that are bold and italic, are SARS-like viruses):
Concluding remarks
Concluding remarks
I generated Z-curves of 39 available genetic sequence of COVID-19 viruses together with 40 SARS-like viruses. An eyeballing of 3D visualization of Z-curves suggests that there are three SARS-like viruses very close to the virus family of COVID-19. By qualifying the similarity/dissimilarity between Z-curves, I identify those three viruses as Bat coronavirus isolate RaTG13 (MN996532), Bat SARS-like coronavirus isolate bat-SL-CoVZXC21 (MG772934 .1), and Bat SARS-like coronavirus isolate bat-SL-CoVZC45 (MG772933 .1). The hierarchical clustering of viruses based on the Z-curves also confirms the visualization, and identify Bat coronavirus isolate RaTG13 as the most-likely culprit of COVID-19. My results strongly support the hypothesis of a Bat origin of COVID-19.
References
References
R. Zhang and C-T Zhang, A Brief Review: The Z-curve Theory and its Application in Genome Analysis, Curr Genomics. 2014 Apr; 15(2): 78–94.
W-X Zheng et al, Coronavirus phylogeny based on a geometric approach, Molecular Phylogenetics and Evolution 36 (2005) 224–232